


Description
Malonyl-CoA decarboxylase deficiency is a condition that prevents the body from converting certain fats into energy.
The signs and symptoms of malonyl-CoA decarboxylase deficiency can begin in infancy or childhood and often include:
- Delayed development
- Decreased muscle tone (hypotonia)
- Low blood glucose (hypoglycemia)
- Weakened and enlarged heart muscle (cardiomyopathy)
- Brain abnormalities
- Seizures
- Diarrhea
- Vomiting
- A buildup of a chemical called lactic acid in the body (lactic acidosis)
- Elevated levels of acid in the blood (metabolic acidosis)
- Elevated blood levels of ketones, which are produced during the breakdown of fats, in the blood (ketosis)
If malonyl-CoA decarboxylase deficiency is diagnosed during newborn screening, early treatment may help prevent or lessen many of the serious health problems that are associated with this condition.
Frequency
Malonyl-CoA decarboxylase deficiency is very rare; only about 60 cases have been reported in the scientific literature.
Causes
Genetic changes that cause disease are called pathogenic variants. Pathogenic variants in the MLYCD gene cause malonyl-CoA decarboxylase deficiency. The MLYCD gene provides instructions for making an enzyme called malonyl-CoA decarboxylase. Malonyl-CoA decarboxylase is responsible for the chemical reaction that converts a molecule called malonyl-CoA into a molecule called acetyl-CoA, which is then used to make new fatty acids. Many tissues, including the heart and brain, use fatty acids as a major source of energy. When malonyl-CoA decarboxylase is not active, malonyl-CoA stops other enzymes from breaking down fatty acids.
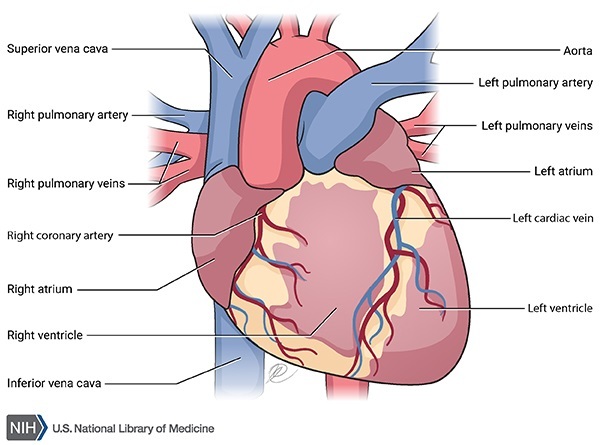
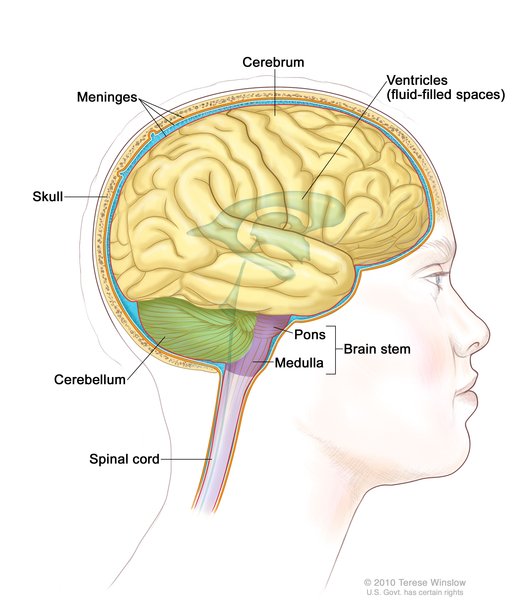
Pathogenic variants in the MLYCD gene prevent malonyl-CoA decarboxylase from functioning properly. A shortage of functional malonyl-CoA decarboxylase disrupts the normal balance of fatty acid formation and breakdown in the body. Without enough functional enzyme, malonyl-CoA cannot be converted to acetyl-CoA, which leads to a buildup of malonyl-CoA. When too much malonyl-CoA is present, fatty acids are not converted to energy. This seems to have a particular impact on the brain and heart, although it is unclear how the specific features of malonyl-CoA decarboxylase deficiency, such as cardiomyopathy and developmental delays, occur.
Inheritance
Malonyl-CoA decarboxylase deficiency is inherited in an autosomal recessive pattern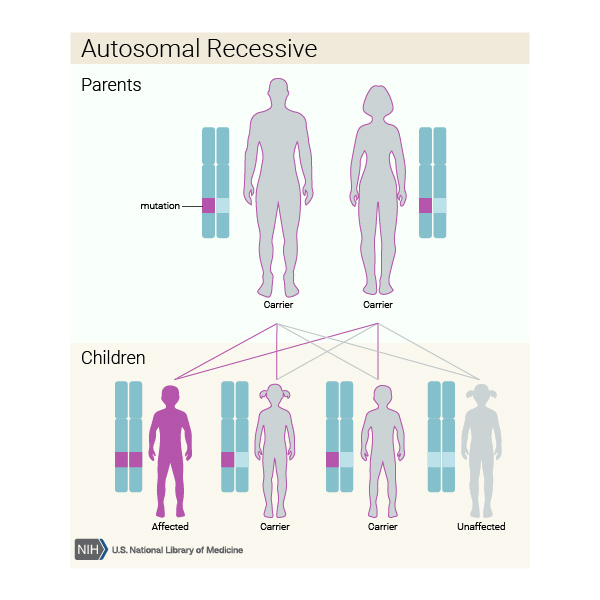 , which means both copies of the gene in each cell must have a pathogenic variant to cause the disorder. The parents of an individual with an autosomal recessive condition each carry one copy of the altered gene, but they typically do not show signs and symptoms of the condition.
Other Names for This Condition
- Deficiency of malonyl-CoA decarboxylase
- Malonic acidemia
- Malonic aciduria
- Malonyl-coenzyme A decarboxylase deficiency
- MCD deficiency
- MYLCDD
Additional Information & Resources
Genetic Testing Information

Genetic and Rare Diseases Information Center
Patient Support and Advocacy Resources
Clinical Trials
Catalog of Genes and Diseases from OMIM
Scientific Articles on PubMed
References
- Chapel-Crespo C, Gavrilov D, Sowa M, Myers J, Day-Salvatore DL, Lynn H, Regier D, Starin D, Steenari M, Schoonderwoerd K, Abdenur JE. Clinical, biochemical and molecular characteristics of malonyl-CoA decarboxylase deficiency and long-term follow-up of nine patients. Mol Genet Metab. 2019 Sep-Oct;128(1-2):113-121. doi: 10.1016/j.ymgme.2019.07.015. Epub 2019 Jul 29. No abstract available. Citation on PubMed
- Kasapkara CS, Civelek Urey B, Ceylan AC, Unal Uzun O, Cetin II. Malonyl coenzyme A decarboxylase deficiency with a novel mutation. Cardiol Young. 2021 Sep;31(9):1535-1537. doi: 10.1017/S104795112100113X. Epub 2021 Mar 22. Citation on PubMed
- Salomons GS, Jakobs C, Pope LL, Errami A, Potter M, Nowaczyk M, Olpin S, Manning N, Raiman JA, Slade T, Champion MP, Peck D, Gavrilov D, Hillman R, Hoganson GE, Donaldson K, Shield JP, Ketteridge D, Wasserstein M, Gibson KM. Clinical, enzymatic and molecular characterization of nine new patients with malonyl-coenzyme A decarboxylase deficiency. J Inherit Metab Dis. 2007 Feb;30(1):23-8. doi: 10.1007/s10545-006-0514-6. Epub 2006 Dec 20. Citation on PubMed
- Zhang JM, Hao LL, Qiu WJ, Zhang HW, Chen T, Ji WJ, Zhang Y, Liu F, Gu XF, Yang SH, Han LS. Clinical, biochemical and genetic characteristics and long-term follow-up of five patients with malonyl-CoA decarboxylase deficiency. Brain Dev. 2024 Oct;46(9):286-293. doi: 10.1016/j.braindev.2024.07.001. Epub 2024 Jul 27. Citation on PubMed
- Zhao C, Peng H, Jiang N, Liu Y, Chen Y, Liu J, Guo Q, Wu Z, Wang L. A case of malonyl coenzyme A decarboxylase deficiency with novel mutations and literature review. Front Pediatr. 2023 Apr 17;11:1133134. doi: 10.3389/fped.2023.1133134. eCollection 2023. Citation on PubMed
The information on this site should not be used as a substitute for professional medical care or advice. Contact a health care provider if you have questions about your health.